EU Clinical Trials Regulation & CTIS
All EU clinical trials now run through CTIS, with built-in transparency rules that make trial information and documents public according to defined publication rules and timelines.
Last verified October 2026 against primary sources.
At a glance
- Authority
- European Medicines Agency, which operates CTIS. Each EU/EEA Member State’s competent authority and ethics committee assesses applications and enforces the Regulation nationally.
- Legal basis
- Regulation (EU) No 536/2014 (Clinical Trials Regulation), plus the revised CTIS transparency rules (EMA/263067/2023), adopted 5 October 2023
- Applies to
- All interventional clinical trials of medicinal products conducted in the EU/EEA — including non-EU sponsors, who must appoint an EU legal representative (Article 74)
- Trigger
- Running a trial in the EU/EEA requires prior authorization via CTIS; the results clocks start at the end of the trial in all Member States concerned
- Deadlines
- Summary of results + lay summary: 12 months after trial end, 6 months for trials that include pediatric participants; clinical study report: 30 days after the marketing authorisation decision
- Publication
- Public by default under Article 81(4); deferrals removed as of 18 June 2024 — only Category 1 (phase I-type) adult trial results wait 30 months after trial end
- Enforcement
- Member State-level penalties — Article 94 mandates “effective, proportionate and dissuasive” penalties, expressly covering failure to submit information intended for publication
- Status & key dates
- Fully in force: applicable 31 January 2022; CTIS mandatory for new applications since 31 January 2023; all ongoing trials under the CTR since 31 January 2025
Who this is for
Trial managers working through EU CTIS.
The CTIS system turns one submission into a schedule of dated obligations. Each becomes a task with a date derived from your study, not re-typed from a guidance document.
Deadlines that matter
| Event | Deadline | Notes |
|---|---|---|
| Regulation (EU) 536/2014 becomes applicable; CTIS goes live | 31 January 2022 | Replaced the Clinical Trials Directive 2001/20/EC; for one year sponsors could still choose to apply under the Directive. |
| CTIS mandatory for all new clinical trial applications | 31 January 2023 | Every new EU/EEA trial application must be submitted through CTIS — no more Directive-route applications. |
| Directive-era trials transition to the CTR | 31 January 2025 | Any trial approved under Directive 2001/20/EC still running must comply with the CTR and have its information recorded in CTIS. |
| End-of-trial notification | 15 days from trial end | Article 37(1)-(3) sets three separate notifications through the EU portal, each within 15 days: end of the trial in a single Member State, end of the trial in all Member States concerned, and global end of the trial. |
| Summary of results + lay summary | 12 months from end of trial in all Member States concerned | Article 37(4); content per Annexes IV and V. Published on submission, except Category 1 adult trials. |
| Results for trials including pediatric participants | 6 months from trial end | Per EMA CTIS guidance (EMA/441151/2024): trials including pediatric participants submit within 6 months of trial completion. Applies to both the results summary and the lay summary. |
| Results of a protocol-specified intermediate data analysis | 1 year from the intermediate analysis date | Article 37(8) — a separate clock that runs before the end of the trial. |
| Clinical study report submission | 30 days after the marketing authorisation decision | Article 37(4): due after the MA is granted, the procedure completes, or the application is withdrawn. Published upon submission. |
| Revised CTIS transparency rules apply | 18 June 2024 | Deferral mechanism (up to 7 years) removed; CCI protection now works through redaction. New CTIS public portal launched the same day. |
| Category 1 (phase I-type) adult trial results publication | 30 months after end of trial in EU/EEA | An EMA publication clock, not a sponsor action — but it defines when competitively sensitive early-phase results become public. |
Regulatory guidance changes. Deadlines and requirements shown here are a summary, not legal advice — confirm them against the current text published by the relevant authority before you rely on them for a submission.
When is your summary of results due?
- Summary of results + lay summary due Standard trials — CTIS posting deadline +12 months
- Pediatric trials Shortened clock for trials in minors +6 months
Informational only — confirm against the regulation and your regulatory counsel.
Don’t let these dates slip. We’ll email them to you with a calendar file that reminds you 90, 30 and 7 days before each one.
Regulatory guidance changes. Deadlines and requirements shown here are a summary, not legal advice — confirm them against the current text published by the relevant authority before you rely on them for a submission.
EU Clinical Trials Regulation disclosure workflow
-
Map every trial to its CTIS category and clocks.
Inventory all EU/EEA trials, assign each to transparency Category 1 (phase I, bioequivalence and biosimilarity), Category 2 (phase II–III), or Category 3 (phase IV and low-intervention), and log end-of-trial dates. Those dates drive the 15-day notifications and the 12-month (or 6-month pediatric) results deadlines.
-
Author documents for publication, not just submission.
Since 18 June 2024 the revised CTIS transparency rules have eliminated the prior deferral mechanisms — protocols and other documents publish according to trial category and document type. Write with EU Clinical Trials Regulation disclosure in mind and plan redaction of personal data and CCI before anything is uploaded.
-
Stand up the lay summary workflow early.
The layperson summary is due with the results summary — 12 months after trial end, 6 for pediatric — and EU guidance (Good Lay Summary Practice) expects translations into the local languages of each Member State concerned. Start drafting at database lock, not at month eleven.
-
Run the notification and results calendar.
The clinical trial regulation EU sponsors answer to runs on dates: the 15-day end-of-trial notifications, the results submission clocks, intermediate-analysis clocks under Article 37(8), and the 30-day CSR window where the trial was intended to support a marketing authorisation.
-
Document your redaction and CCI rationale.
Article 81(4) allows confidentiality only on specific grounds. Keep a defensible record of what was redacted, why it qualifies as CCI or personal data, and the anonymization method — particularly for documents subject to publication through CTIS.
EU CTR / CTIS vs FDAAA 801 (ClinicalTrials.gov)
For sponsors running trials on both sides of the Atlantic — differences only. The EU CTR requirement itself is in At a glance above.
| Dimension | EU CTR / CTIS | FDAAA 801 / ClinicalTrials.gov |
|---|---|---|
| Scope | All interventional clinical trials of medicinal products conducted in the EU/EEA, Phase 1 included. | Applicable clinical trials of FDA-regulated drugs, biologics and devices with a US nexus (42 CFR Part 11) — Phase 1 drug trials and small feasibility device studies are excluded. |
| Registration trigger | Authorization is required before the trial may start — application through CTIS, mandatory for new applications since 31 January 2023. | Register within 21 calendar days after the first participant is enrolled (42 CFR 11.24). |
| Results clock | 12 months after end of trial in all Member States concerned; 6 months where pediatric participants are included. | 12 months after the primary completion date (42 CFR 11.44). |
| What is published | Structured data plus documents: protocol and application documents on the category timing, the summary of results, the layperson summary, and the CSR after a marketing authorisation decision. | The registration record, structured tabular results — participant flow, baseline characteristics, outcome measures and adverse events — and the full protocol and statistical analysis plan. No patient-level data. |
| Document redaction | The sponsor redacts personal data and CCI from documents before upload, and personal-data protection applies to published documents at all times. | The protocol and SAP go up with the results, and personal and confidential details can be redacted first (42 CFR 11.48), the same document work ANONYMIZE handles. |
| Delay options | None since 18 June 2024 — the deferral mechanism was removed. The only built-in lag is the 30-month publication timing for Category 1 adult trial results. | A certification for an unapproved or uncleared product delays results to as late as 2 years after certification, and good-cause extensions are available (42 CFR 11.44(b), (e)). |
| Enforcement | Set by each Member State under Article 94, which requires penalties that are “effective, proportionate and dissuasive” — there is no EU-wide fixed fine. | FDA pre-notices, notices of noncompliance and civil money penalties of up to $10,000, plus up to $10,000 for every day a violation stays uncorrected after FDA’s notice: $15,107 a day with the 2025 inflation adjustment (21 U.S.C. 333(f)(3)). |
Where they agree Both put the obligation on the sponsor as responsible party, and both results clocks run from the trial’s own completion rather than from the publication of any paper — though the registration triggers differ, as the second row shows.
Self-assessment
How ready is your disclosure program?
Eight questions, two minutes, no form until the end. You get a readiness score by area and the three things to fix first. Email is only asked for if you want the full breakdown sent to you.
Want the full breakdown?
We’ll email your area-by-area scores with the specific CTIS rules behind each question, and what a fix looks like at your scale. Work email, no newsletter unless you ask.
Regulatory guidance changes. Deadlines and requirements shown here are a summary, not legal advice — confirm them against the current text published by the relevant authority before you rely on them for a submission.
Last reviewed: October 1, 2026
What is the EU Clinical Trials Regulation (EU CTR) and CTIS?
The EU CTR is Regulation (EU) No 536/2014, the law governing interventional clinical trials of medicines in the EU/EEA, and CTIS is the Clinical Trials Information System, the single submission portal and public database it created. Applicable since 31 January 2022, the regulation makes trial information and documents public under defined transparency rules and puts results summaries, including lay summaries, on hard 12-month and 6-month clocks.
What the EU CTR requires
Sponsors must obtain authorization through CTIS before a trial may start. Article 37(1)-(3) then requires notification through the EU portal within 15 days of the end of the trial.
Under Article 37(4), a summary of results and a plain language summary (PLS) — the regulation’s summary written for laypersons — follow in the content set out in Annexes IV and V, within 12 months of the end of the trial in all Member States concerned, or 6 months for trials that include pediatric participants; a clinical study report is due within 30 days when the trial was intended to support a marketing authorisation, following the relevant marketing authorisation decision, completion of the procedure, or withdrawal of the application. Personal data and commercially confidential information must be handled in the document versions before upload, and a sponsor not established in the Union must appoint an EU legal representative (Article 74).
What that means operationally
- Disclosure decisions embedded in trial conduct, not bolted on at the end.
- Document versions prepared for publication — protocols and reports with personal data and CCI handled before upload.
- A PLS per trial, in local languages, on a hard clock.
- Ongoing CTIS maintenance: updates, notifications, deadline tracking.
How the platform answers it
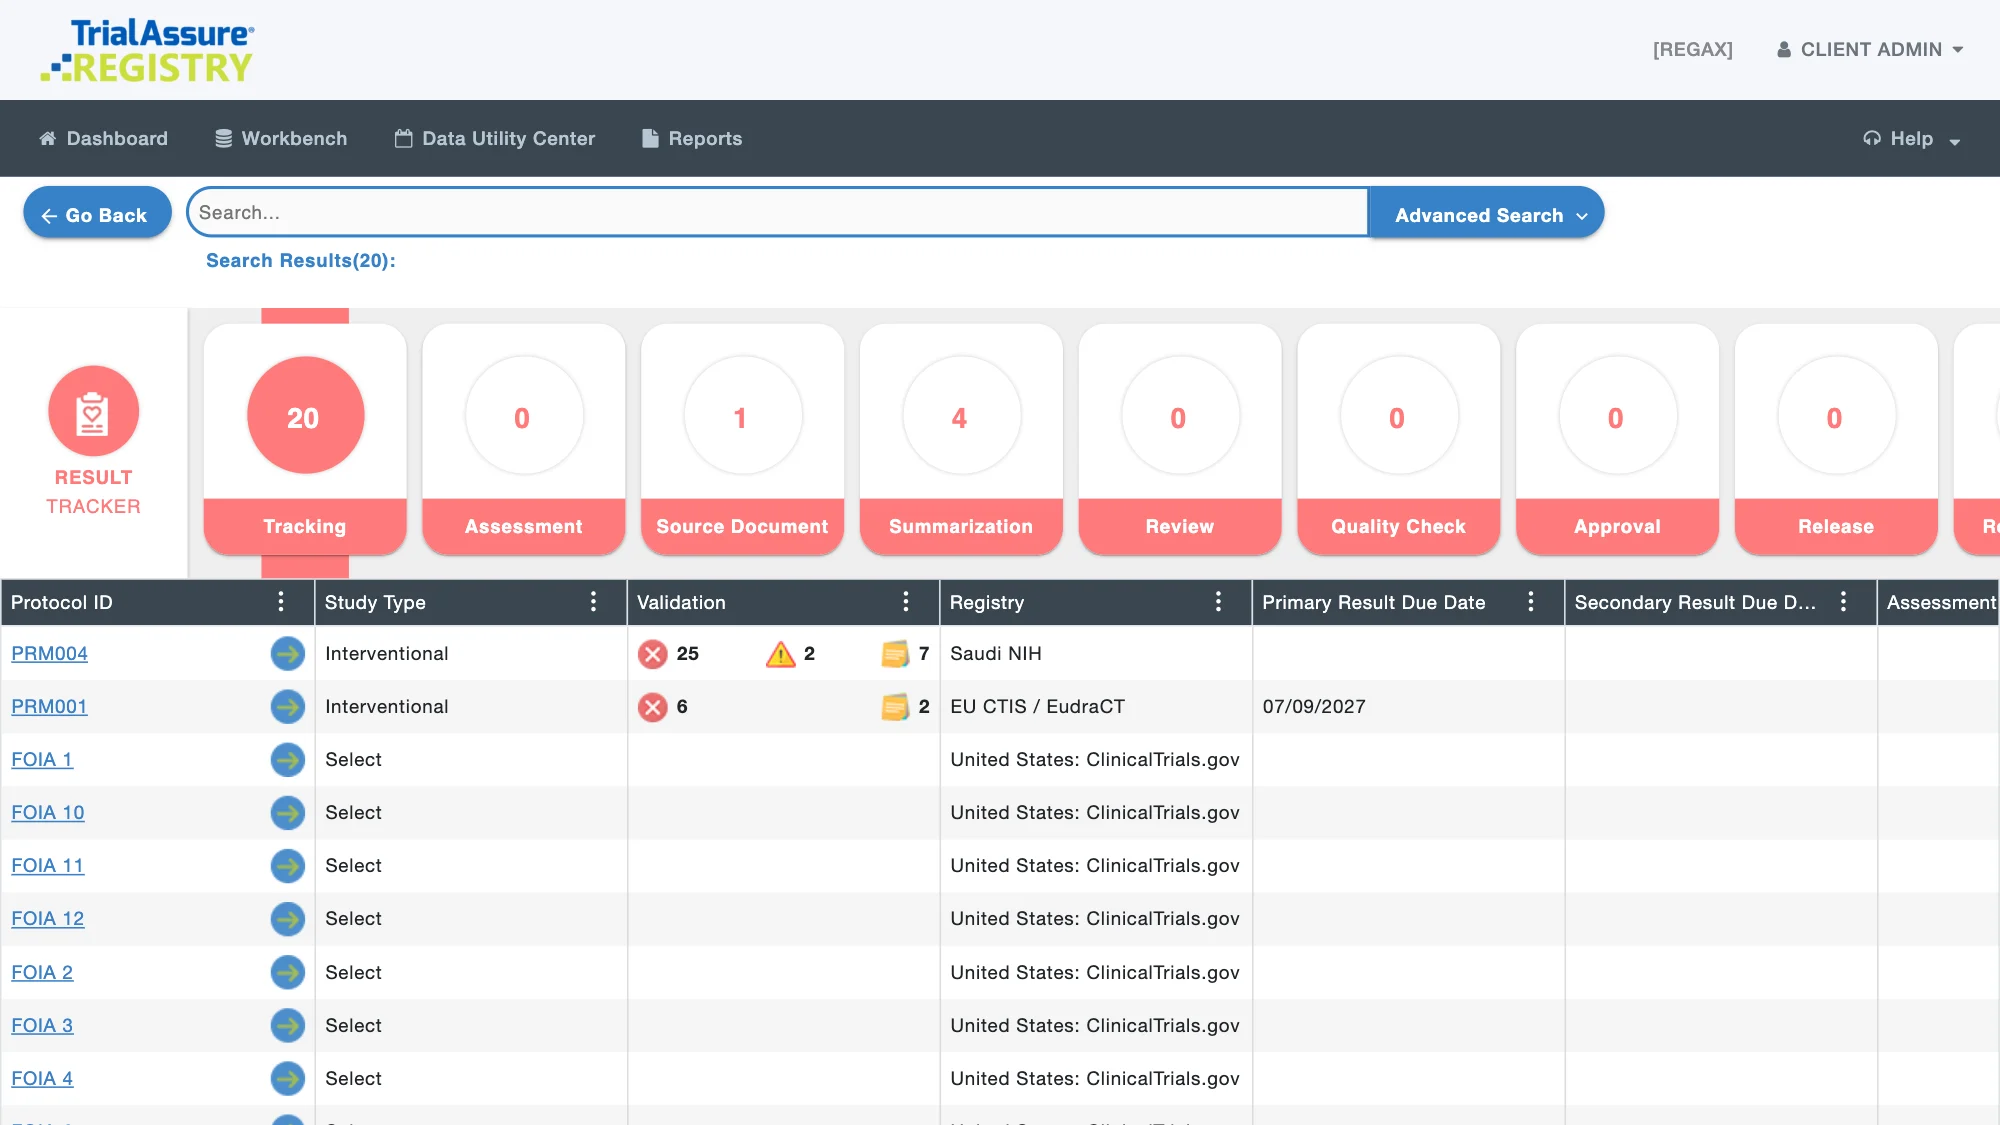
REGISTRY manages CTIS alongside your other 40+ registries. ANONYMIZE prepares publication-ready document versions. LINK AI drafts the PLS — the workflow one pharma sponsor runs, reporting up to 80% faster delivery with medical writers remaining in control of review and finalization. The rules engine tracks the applicable results-submission clocks. EU delivery via TrialAssure B.V.
EU CTR / CTIS — frequently asked questions
What are the EU CTR transparency requirements?
The EU CTR transparency requirements make trial information and the key documents submitted through CTIS public by default. Article 81(4) allows confidentiality only on specific grounds, so the application documents including the protocol, the summary of results, the layperson summary, and the clinical study report after a marketing authorisation decision all reach the public portal. When each item appears depends on the trial’s transparency category — Category 1 (phase I, bioequivalence and biosimilarity), Category 2 (phase II–III) or Category 3 (phase IV and low-intervention) — and since 18 June 2024 there is no deferral mechanism to push it back. What sponsors control is content: personal data and commercially confidential information must be handled before upload.
What is the EU CTR lay summary requirement?
The EU CTR requires a summary of results written to be understandable to laypersons — the regulation’s term for a plain language summary — within 12 months of the end of the trial in all Member States concerned, or 6 months for trials including pediatric participants. It is due alongside the technical results summary, follows the content set out in Annexes IV and V, and EU guidance (Good Lay Summary Practice) expects translation into the local languages of each participating Member State. It publishes on submission, except for Category 1 adult trials, so drafting should start at database lock rather than month eleven.
Can TrialAssure® prepare CTIS document versions?
Yes. TrialAssure ANONYMIZE prepares the publication versions of protocols, reports and appendices — removing or masking personal data and commercially confidential information — and produces the risk documentation that explains and defends the method behind each decision. That record matters because Article 81(4) permits confidentiality only on specific grounds, and Member States can challenge what was withheld. Every transformation is logged and attributable, and our specialists review the output before it goes back to you: AI Enabled. Human Driven.™ Because deferrals ended on 18 June 2024, this work belongs before upload, not after.
Do you handle PLS translations?
Yes — translation is part of the plain language summary service, not a separate hand-off. EU guidance expects the layperson summary in the local languages of each Member State concerned, so we treat the source-language draft and its translations as one deliverable on one clock. LINK AI drafts the summary inside Microsoft Word on your own templates, with per-statement source references back to the results, and our specialists review the source version and each translation before submission. It is the workflow Daiichi Sankyo runs at up to 80% faster. See the Plain Language Summaries page.
Does the EU CTR apply to trials approved under the old Clinical Trials Directive?
Yes — since 31 January 2025, every trial still running in the EU/EEA must comply with Regulation (EU) 536/2014, including trials originally approved under Directive 2001/20/EC. Sponsors had a three-year transition window, from 31 January 2022 to 31 January 2025, to move ongoing trials into CTIS. A legacy Directive approval no longer exempts a trial from the clinical trial regulation EU regulators now enforce, including its transparency and results-reporting requirements.
What happens if a sponsor misses an EU CTR disclosure deadline?
Penalties are set at Member State level: Article 94 of Regulation (EU) 536/2014 requires each country to impose “effective, proportionate and dissuasive” penalties, and it expressly names failure to submit information intended for publication in the EU database as an infringement. There is no single EU-wide fine — exposure varies by country. The gap is also publicly visible on the EU CTIS public portal. TrialAssure tracks every CTIS results clock so deadlines surface before they are missed.
Can sponsors still defer publication of documents in CTIS?
No — the deferral mechanism was removed when EMA’s revised transparency rules took effect on 18 June 2024. Sponsors could previously delay publication of certain data and documents for up to seven years after trial end; now documents publish at their category’s milestone, and commercially confidential information is protected through redaction instead. One timing exception remains: results of Category 1 (phase I-type) trials in adults publish 30 months after trial end. TrialAssure ANONYMIZE prepares those redacted, publication-ready document versions.
Does the EU CTR apply to sponsors based outside the EU?
Yes — the regulation applies to any interventional trial of a medicinal product conducted in the EU/EEA, regardless of where the sponsor is headquartered. Article 74 requires a sponsor not established in the Union to appoint an EU-based legal representative responsible for the sponsor’s compliance (Member States may accept a local contact person for trials run solely on their territory). US and other non-EU sponsors carry the same CTIS submission requirements, transparency duties, and results obligations.
What is the difference between EudraCT and CTIS?
EudraCT was the application database for trials authorized under the Clinical Trials Directive (2001/20/EC); CTIS is its replacement under Regulation (EU) 536/2014 and the single entry point for EU trial applications since 31 January 2023. Directive-approved trials still running had to move to the CTR and record their information in CTIS by 31 January 2025. Teams with mixed-era portfolios end up managing disclosure records that span both systems.
Have our specialists run your CTIS clocks and document prep
In the scope box, tell us how many EU/EEA trials you are carrying, which ones are nearing the results and lay summary deadline, and how many documents still need personal data and CCI handled before upload. We come back with who would run it, what that volume takes, and how soon we can start. A disclosure specialist replies within one business day.
Get CTIS-ready without the fire drill.
Trusted by clinical development teams at sponsors and CROs